Computational drug discovery is getting a major overhaul as researchers shift away from opaque deep learning architectures toward simpler, transparent mathematics. In a study published in November 2026 in the journal Molecular Informatics (doi: 10.1002/minf.70060), a cross-institutional research team led by scientists at Louisiana State University and Furman University unveiled a linear alternative called Geometric Manifold Alignment.
Traditional deep learning methods often struggle to generalize when trained on small biological datasets because they rely on complex neural networks. To solve this bottleneck, the researchers introduced the Multi-View Linear Manifold (MVLM) approach. Instead of adding architectural layers, MVLM aligns the latent spaces of existing foundation models—specifically ChemBERTa for small molecules and ESM-2 for protein sequences—into a shared geometric interaction space.
Benchmarking Against Industry Standards
To build and test the model, the team pretrained MVLM on the Universal Foundation Set, a massive repository comprising 473,923 high-confidence interactions. Within this framework, binding affinity is calculated directly through geometric proximity between drug and target embeddings.
How does an embryo reliably "compute" its form – "cell by cell" – using only local interactions and mechanics, yet produce a precise global body plan? I’m excited to share our Nature Methods paper "MultiCell: geometric learning in multicellular development", presenting… pic.twitter.com/bmyjIClvTK
The performance metrics indicate a substantial leap forward for linear modeling in bioinformatics. MVLM achieved an Area Under the Receiver Operating Characteristic (AUROC) score of 0.936 on the KIBA dataset and 0.928 on the Davis dataset. These figures establish a new state-of-the-art benchmark, outperforming existing computational methods with statistical significance (p < 0.001).
Unlike black-box neural networks that obscure how decisions are made, MVLM offers intrinsic geometric interpretability. Interaction scores function simply as cosine similarities within a shared manifold. Remarkably, the model recovered known kinase phylogenetic relationships without any explicit supervision, demonstrating that aligning pretrained representations captures true biological organization.
Linear Versus Network-Based Approaches
While the LSU and Furman researchers focused on geometric alignment of foundation models, alternative linear strategies are tackling drug-target interaction (DTI) prediction from different angles. As reported by pmc.ncbi.nlm.nih.gov in PMC13501308, researchers investigating the Linear Interpretable Drug-Target Interaction (LI-DTI) model draw inspiration from recommender systems.
While MVLM aligns pre-trained chemical and biological embeddings into a joint space, LI-DTI learns directly from heterogeneous drug-drug and target-target similarity matrices. Both approaches reflect a broader industry pushback against heavy deep learning pipelines like DTINet and NeoDTI, which utilize complex graph diffusion and end-to-end matrix completion.
| Model Name | Core Mechanism | Primary Datasets / Benchmarks | Key Advantage |
|---|---|---|---|
| MVLM | Multi-View Linear Manifold geometric alignment | KIBA, Davis, Universal Foundation Set | Recovers kinase phylogeny without supervision; AUROC 0.936 (KIBA) |
| LI-DTI | Linear combination of similarity matrices | DrugBank (2011 to 2022 prospective validation) | Transparent predictions for drugs or targets with no prior known interactions |
Both MVLM and LI-DTI emphasize transparency and practical usability, helping researchers generate robust hypotheses for downstream experimental prioritization.
Worth a look
3 Best Expert-Backed Bets For Suns vs Nuggets Tonight – Sports Illustrated
The Phoenix Suns face the Denver Nuggets tonight at the Mortgage Matchup Center with both teams seeking momentum...

California Runaway and Homeless Youth Grantees
California Deploys Millions Across Dozens of Local Networks to Tackle Youth Homelessness Funding allocations span multiple community organizations...
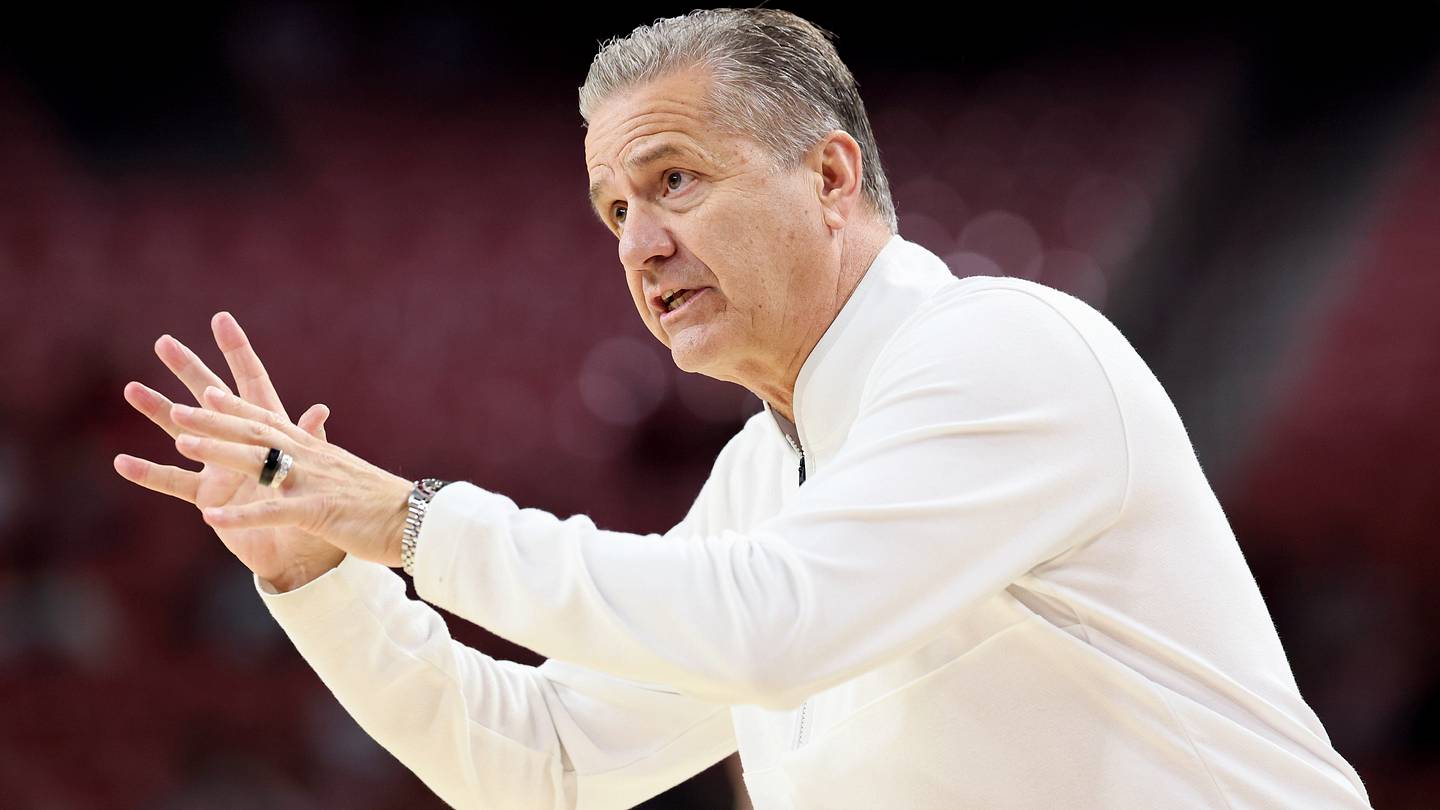
Arkansas Basketball Projected Rotation: Who Starts, Who Backs Up, What’s Unsettled
Calipari Weighs Rotation Choices as Arkansas Basketball Opens Season Against UA-Pine Bluff Fayetteville, Ark. — With the season...
Utah football is, once again, Thunderstruck – College of Social and Behavioral Science
Bringing Back the Thunder: How the Ute Thunder Cannon Returned to Utah Football Back in 2003, the Utah...